Friday, 3 February 2017

CECAM Workshop - Designing Forcefields in an Age of Cheap Computing July 26, 2017 - July 28, 2017

FMD researchers Prof John Harding, Dr Colin Freeman and Dr Chris Handley recently won funding to host a workshop at the University of Sheffield, the topic being "Designing Forcefields in an Age of Cheap Computing", July 26, 2017 - July 28, 2017.

Robust, reliable forcefields are central to successful atomistic simulations. This workshop will bring together leading experts to discuss the impact of increasing computer power, both in terms of speed and data storage, on the development, validation and use of forcefields in molecular simulation.

Chris: As a researcher who has worked in the field of chemical simulation since my master project with Prof Paul Popelier, it is exciting to be able to host a workshop such as this, bringing together experts in atomistic simulations for all manner of materials and molecules, and to discuss the problems that still persist in traditional simulation methods, and how they might be addressed with cutting edge computing techniques. It should lead to lively discussions and some good insight into how such simulations can take advantage of modern computing power.

Invited speakers will also include experts in machine learning and GPU programming, who can give an informed perspective on modern HPC computing.

The workshop by its nature with feature keynote presentations, framing particular topics in chemical and materials simulations, with more open discussion forums between talks, allow groups to break off and tackle topics in more depth.

Short talks and posters are welcome.


Delegates will be hosted at Halifax Hall, located on the west side of Sheffield City Centre and only a short walk from Ecclesall Road, an area which provides a hub of bars, pubs and restaurants. It offers a great location for both business and leisure trips in Sheffield. Situated within The University of Sheffield’s Endcliffe student village it also offers the perfect place for visiting parents and prospective students to stay.

Local attractions include the picturesque Botanical Gardens, only five minutes away, whilst shopping and other city centre attractions are easily commutable by bus and taxi. Further afield the location offers great access to The Peak District National Park, which by car is only a 15 minute drive, whilst the stunning Chatsworth House is only a 30 minutes’ drive.


ICACC - International Conference and Expo on Advanced Ceramics and Composites

The 41st InternationalConference and Expo on Advanced Ceramics and Composites (ICACC) was held on 22-27 January 2017 at Daytona Beach, Florida, USA. This annual conference showcases cutting-edge research in advanced ceramics, armour ceramics, solid oxide fuel cells, ceramic coatings, bio-ceramics and more. Postdoctoral Research Associate Dr. Fan Yang from our group attended the conference and presented our recent work on tuning the electrical properties of sodium bismuth titanate perovskite by chemical doping with a focus on the oxide-ion conduction behaviour of NBT. Her presentation has raised lots of attention from the audience who also work on developing new electrolyte/electrode materials for intermediate temperature solid oxide fuel cells (IT-SOFCs).


Some key invited speakers working on oxide-ion/mixed conductors include Professor Eric Wachsman from University of Maryland, USA, Professor Tatsumi Ishihara from Kyushu University, Japan and Professor Scott Barnett from Northwestern University, USA. Other symposiums related to our group’s work include lithium/sodium ion batteries, thermoelectrics and piezoelectrics, etc.

Hilton Daytona Beach Resort where the conference 
was held and the beautiful sea view outside Hilton


Friday, 20 January 2017

FMD group’s expertise highlighted in new Chemistry of Materials review


The ceramic processing and thermoelectric characterisation expertise in the FMD group has been recently highlighted in an invited review in Chemistry of Materials. The paper, part of a major special edition of the journal, covered the major aspects of lab-based ceramic processing, and the standards required for reliable thermoelectric measurements, and gives a step-by-step guide to making high quality, reproducible ceramics using solid state processing. It also covers the major techniques and potential pitfalls associated with the characterisation of thermoelectric properties in n-type ceramics.

January’s special edition comprises a series of articles from internationally recognised groups, containing protocols and “trade secrets” in a wide variety of research areas, direct from the researchers in the field, and we were very pleased to be included in this wide ranging edition. 


The full reference is as follows: 

Protocols for the Fabrication, Characterization, and Optimization of n-Type Thermoelectric Ceramic Oxides

R. Boston , W. L. Schmidt, G. D. Lewin, A. C. Iyasara, Z. Lu, H. Zhang, D. C. Sinclair, and I. M. Reaney, Chem. Mater. 29 (2017), 265-280. 


Friday, 13 January 2017

N8HPC/CCP5 New Approaches to Atomistic and Quantum Simulation of Materials

Dr David Quigley from the University of Warwick introduces Robyn.
On Friday 6th of January, a couple of us were at the New Approaches to Atomistic and Quantum Simulation of Materials; N8HPC and CCP5 Network Event in York, UK. This one day event brought together computational chemists, solid state materials scientists and more, from the N8 group of universities, plus guests from further afield.

PhD student, Robyn Ward presented her work on the application of metadynamics to explore the migration pathways of ions in BaTiO3. The novelty of her work helps further the understanding of just how the materials work that are made in our ceramic laboratories.


The 5x5x5 supercell of MALI, quenched from 400K to 50K.
Grey lead iodide octahedra show tilting.
Methylammonium ions show ordering in their
alignment not present at room temperatures
Research fellow, Dr Chris Handley also attended, presenting his recently published work on developing a forcefield for Methylammonium Lead Iodide, again another material that is being synthesised within our experimental laboratories.

The event was a great chance to network with those with whom we can share expertise with, and to talk in person with the developers of particular simulation programs. This is particularly useful as we branch out to running new types of simulations and deepening the synergy between our computational and our experimental materials scientists.

Some particular highlights from other speakers at the event was Dr David Quigley from the University of Warwick (where Chris used to work) presenting his work on simulations of Ice 0, and the work of Prof Gabor Csanyi from the University of Cambridge and his machine learning based simulations for materials. 


Wednesday, 21 December 2016

Vivas and News

In the last couple of week, Fayaz, Simon, and Amir have all passed their PhD vivas, and all only need to submit minor corrections. Congratulations to all of them.


This of course means all three, along with Leo, are leaving our group for pastures new and we wish them the best of luck in their future endeavours.

John Harding, Colin Freeman, and Chris Handley also received news that they had obtained funding for a CECAM workshop that they will be hosting in Sheffield next summer. The workshop - Designing Forcefields in an Age of Cheap Computing - will bring together experts from different fields of atomistic simulation to address the challenges of chemical and materials simulations in the coming years.

On the 6th of January, Chris Handley, and PhD student Robyn Ward, will be giving presentations at the N8 / CCP5 one day conference, New Approaches to Atomistic and Quantum Simulation of Materials. Chris will be talking about his work on methylammonium lead iodide, while Robyn will be talking about her work on barium titanate.

Thursday, 3 November 2016

Bashing Down Windows for Material Science

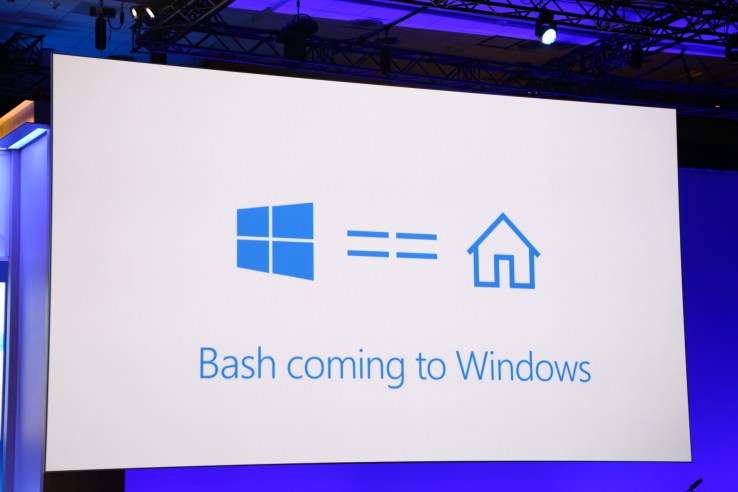
In the last few months Windows 10 has had an interesting new capability – Bash. Originally the Linux Subsystem was only available for those on the developer loop, but since the Windows 10 Anniversary edition this subsystem has been available to all users who activate it. The subsystem is not an emulator, but a way for Windows 10 to run Linux applications, and to use the Linux Bash environment, through the use of dynamic maps between Linux system calls and Windows ones.
As a computational chemist working in the Department of Materials Science and Engineering this is really an excellent and exciting new way that Windows has evolved. There are a great many tools for my research. Some work on Windows, and are well designed for that OS, given that they are applications aimed at the people that make and analyse their materials. These tools help users visualize crystal structures in 3D, or predict from crystal structures experimental observables, such as transition electron microscopy. For computational chemists, these tools are often also invaluable as they allow us to construct visually the crystal structures that we wish to then simulate using quantum mechanics or classical force fields. More often than not, the programs designed for running such chemical simulations, have no GUI, and run in a Unix environment. CASTEP is a UK created Density Functional Theory simulation package, which is free for all UK academics, and is used extensively by those researchers wishing to simulated solid state materials, such as batteries, piezoelectric materials, and solar power materials. Previously, to run CASTEP on a Windows machine, Cygwin or a virtual machine were required. However, with the new subsystem, CASTEP installs out of the box as if you were running any other Linux computer. The same is equally true of GULP, another program used in materials science, which is often used to design, test, and analyse atomistic potentials. DL_POLY, another UK created simulation package is also used by a large user base to perform molecular dynamics simulations using atomistic potentials.
All of the above programs mentioned, and many more, such as the DFT codes VASP and WIEN2K, and other molecular dynamics programs such as GROMACS, and LAMMPS, can have their output analysed by these Windows 10 packages, and their inputs easily designed by these same crystal analysis programs, but natively are best run in a Unix environment.
The typical work around has always been either the use a virtual machine, Cygwin, or, using more expensive Apple computers, or making users use Linux machines for which they may not be comfortable using – especially if their previous workflow used packages that ran on Windows.
Personally I fall into that last category of users. While I can write a paper in LaTeX, I really don’t like it compared to the WYSIWYG world of Word, and of course with word I can use my favourite citation manager, Zotero (which by the way the work around using Dropbox is also good fun). That impact on workflow is an important thing, especially if you are dealing with final year students who you want to work on your research. Ideally you want to get them up and running ASAP where the only teaching you need to do is how to run the simulation packages. I don’t want to have to teach them how to use and entirely new OS, and in the case of Linux, perhaps entirely new ways to write documents and make spreadsheets. This is especially true if the university course from the first year onwards has included access to MS Office, and has done teaching using those tools.
By being able to now run many of these simulation packages through the Windows Bash Linux subsystem there are minimal hoops to jump through. All students now have easy access to a machine that can run the simulation programs, and without having to switch OS, or log into a dedicated UNIX server which is maintained for PhD and postdoc research. That lack of need to use a virtual machine, or emulator, also means much less impact on resources on personal machines, and less peculiarities with the allocation of computing resources on those machine. Furthermore, with respect to workflows, inputs and outputs from those simulation packages all can happen under the one roof of the Windows 10 OS, and lead to greater productivity.
Bash in Windows 10 has trampled down a barrier which makes the use of the OS far more competitive, cost effective and productive for computational chemistry.
You can learn more about research software and the sustainability of such research at the University of Sheffield Research Software and Engineering blog. 

Saturday, 10 September 2016

FMD at ICAMM 2016

ICAMM 2016 is the International Conference for Advanced Materials Modelling, held at the end of August in Rennes, France. The conference was also preceded by a training workshop for the quantum mechanics program, VASP. From the FMD research groups, Dr Chris Handley, and PhD students, Emilio Pradal Velazquez and Stavrina Dimosthenous attended for this 9 day long event. It was a great opportunity to both learn how to use a new computer program for materials discovery, and also meet and watch talks from various experts in different fields of materials design.

A significant number of talks were devoted to the topic of hybrid halide-organic perovskites, and their potential use a photovoltaic material. Chris presented his work (just submitted) on designing a new forcefield for modelling methylammonium lead iodide, which was well received given it allows for far larger simulations of the materials that are not possible when using quantum mechanical simulations.

"With the University of Sheffield bringing on line in the New Year a new HPC server, that makes use of the new generation of GPUs, it will be possible to use VASP to perform significant calculations, in conjunction with NMR and XRD, to understand materials and their properties" - Chris Handley


groupe_2_small.jpeg

You can just see Chris in the front row, 4th in front the right, with Emilio to the left, and Stavrina to Emilio's left.


Part way through the workshop/conference there was an excursion to Mont Saint Michel.